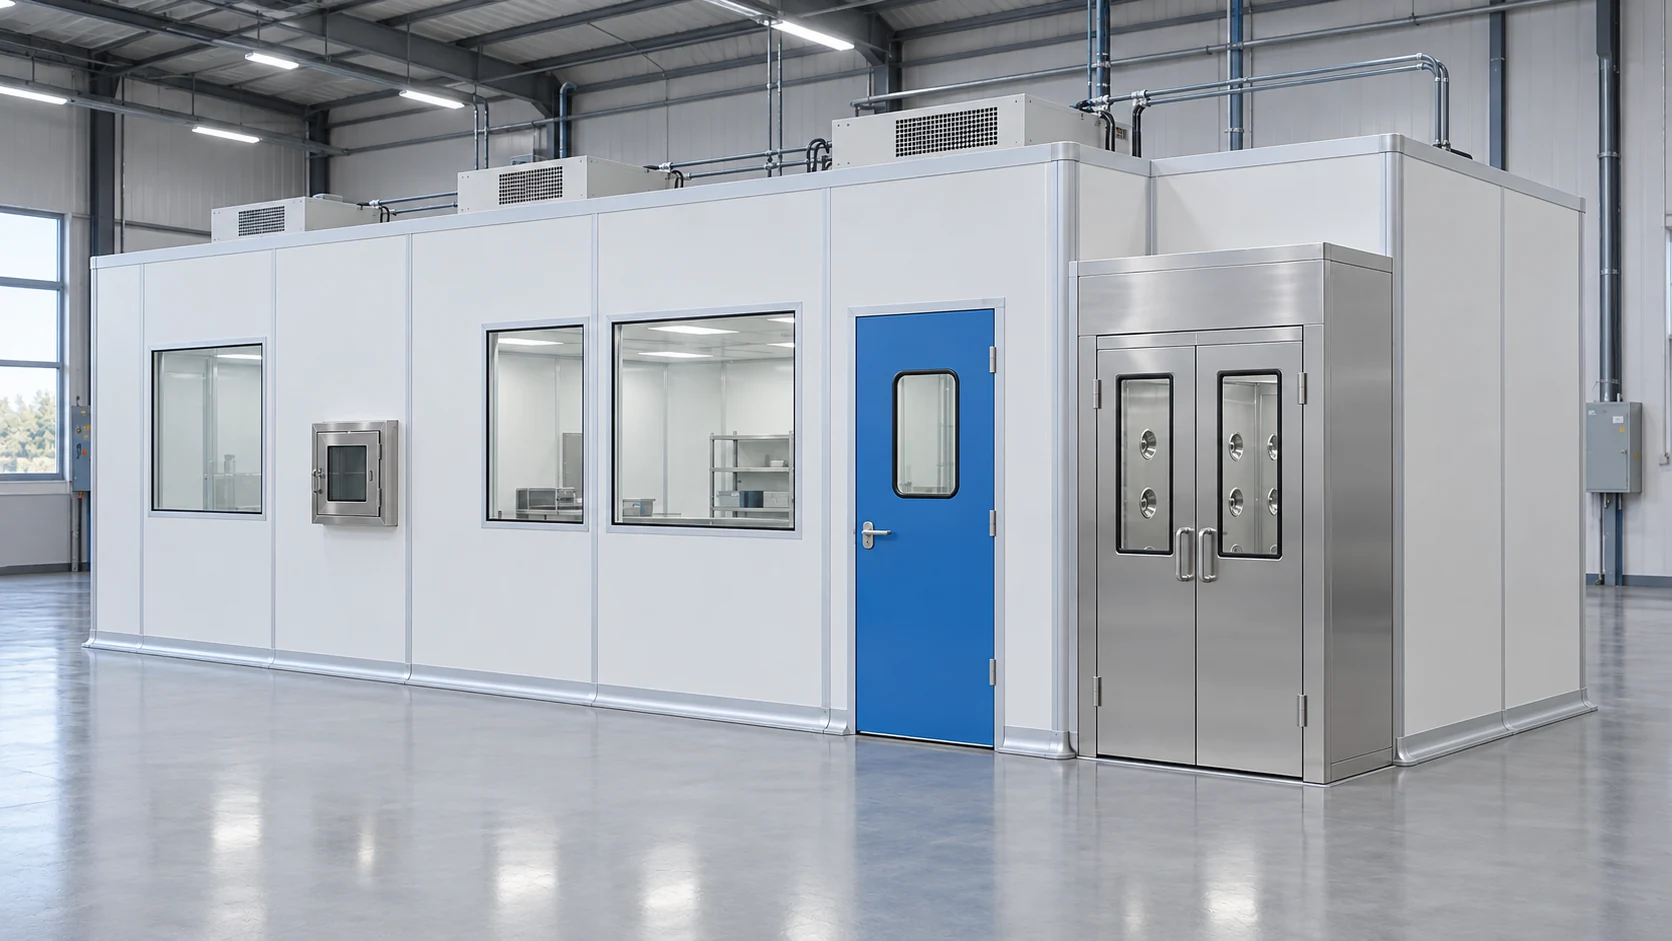
Connect the facility package to the regulated process
Identify the product and operation before setting environmental requirements. Aseptic manufacture of sterile drugs has a different context from many non-sterile or device-related activities. The relevant facility brief must come from the actual process and applicable requirements.
For sterile drug and biological products made by aseptic processing, FDA publishes guidance addressing the CGMP context. Use the appropriate guidance with the quality and facility teams to define the design inputs and required evidence; do not turn its title into a product certification claim.
In procurement, separate the room package from the owner's qualification, operating and monitoring activities. Ask for the drawings, equipment information and agreed test records that support those activities. Keep responsibility for each deliverable explicit.
Configuration decisions
| Decision | What to define |
|---|---|
| Intended process | Define the product, exposed operations and contamination risks. |
| Facility brief | Translate applicable requirements into reviewed design inputs. |
| Supply evidence | Agree drawings, component records and the required acceptance scope. |
| Operational ownership | Assign qualification, procedures and monitoring responsibilities. |
Is there a universal FDA-approved modular room?
A room package should not be presented as a universal regulatory approval. Its suitability depends on the intended operation, installed configuration and the wider quality and compliance framework.
Technical references
Requirements depend on the intended use and project conditions. Consult the relevant current documents when defining the final specification.